Metalla-aromatics have been defined as metallacycles that are derived from the formal replacement of a carbon atom in the framework of an organic aromatic ring with a metal fragment.[1]. Here I explore whether this simple definition can be broadened to sulfur-nitrogen rings which contain few[2],[3] if indeed any[4],[5],[6],[7] carbon atoms and which also happen to be planar aromatic molecules.
Prologue
Our story starts with the authors of this article[8] reporting the formation of the sulfur-nitrogen titanacycles 1[9] and 2[10] (green in the scheme 1 below) from Cp2Ti(CO)2 reacting with S4N4. Isolating compound 2 was something of a surprise, since they had been expecting/hoping to get 3[11] instead (red in scheme 3 below). This result was unexplained in the article and unremarked upon by anyone else since.
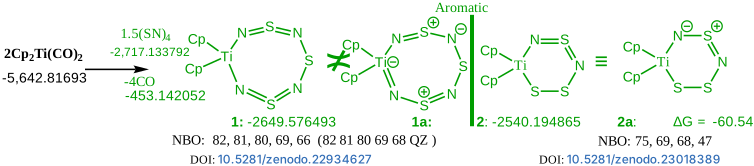
Scheme 1. Formed products 1 – 2 from the reaction of Cp2Ti(CO)2 with S4N4.
To develop the prologue, we start with compound 7 (Scheme 2) which was described[3] as a (Hückel-class) 10 π-aromatic ring (6e from double bonds, 4e from S lone pairs) containing one ring carbon atom and represented in the original paper as 7a. In fact, NBO7 analysis[12],[13]‡ (as obtained with no 3-centre bonds allowed) suggests that 7b is a more accurate representation (4e from two π-double bonds and 6e from lone pairs = 10π =4n+2, n=2 ). Molecule 7b has a computed NICS(0) (= Nucleus Independent Chemical Shift at ring centroid) value of -9.6 ppm,[14] NICS here being used as an approximate indicator of aromaticity (or lack of it), with e.g. the archetypal 4n+2 (n=1) π-aromatic benzene giving a NICS(0) value of ~-10. For 4n/4n+2 aromaticity selection rules, see e.g. [15],[16]
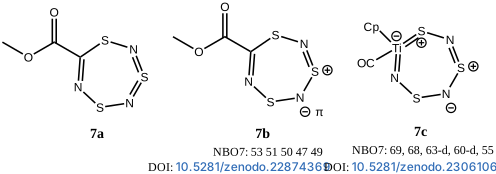
Scheme 2. Compounds 7a/b and 7c.
A replacement of the carbon and its attached ester group according to the definition above by using (Cp.CO)Ti as a transition metal results in e.g. 7c – a compound not dissimilar to the compound discussed in the previous post[17]. A MN15L/Def2-TZVPP calculation[18] gives the geometry shown in Figure 1. A NBO7[12] localisation procedure‡ gives the result[19] shown as 7c (Scheme 2, Figure 2), with back bonding into titanium d-orbitals and the three π-bonds and two lone pairs (one N, one S) resulting in a total of ten cyclically conjugated π-electrons.
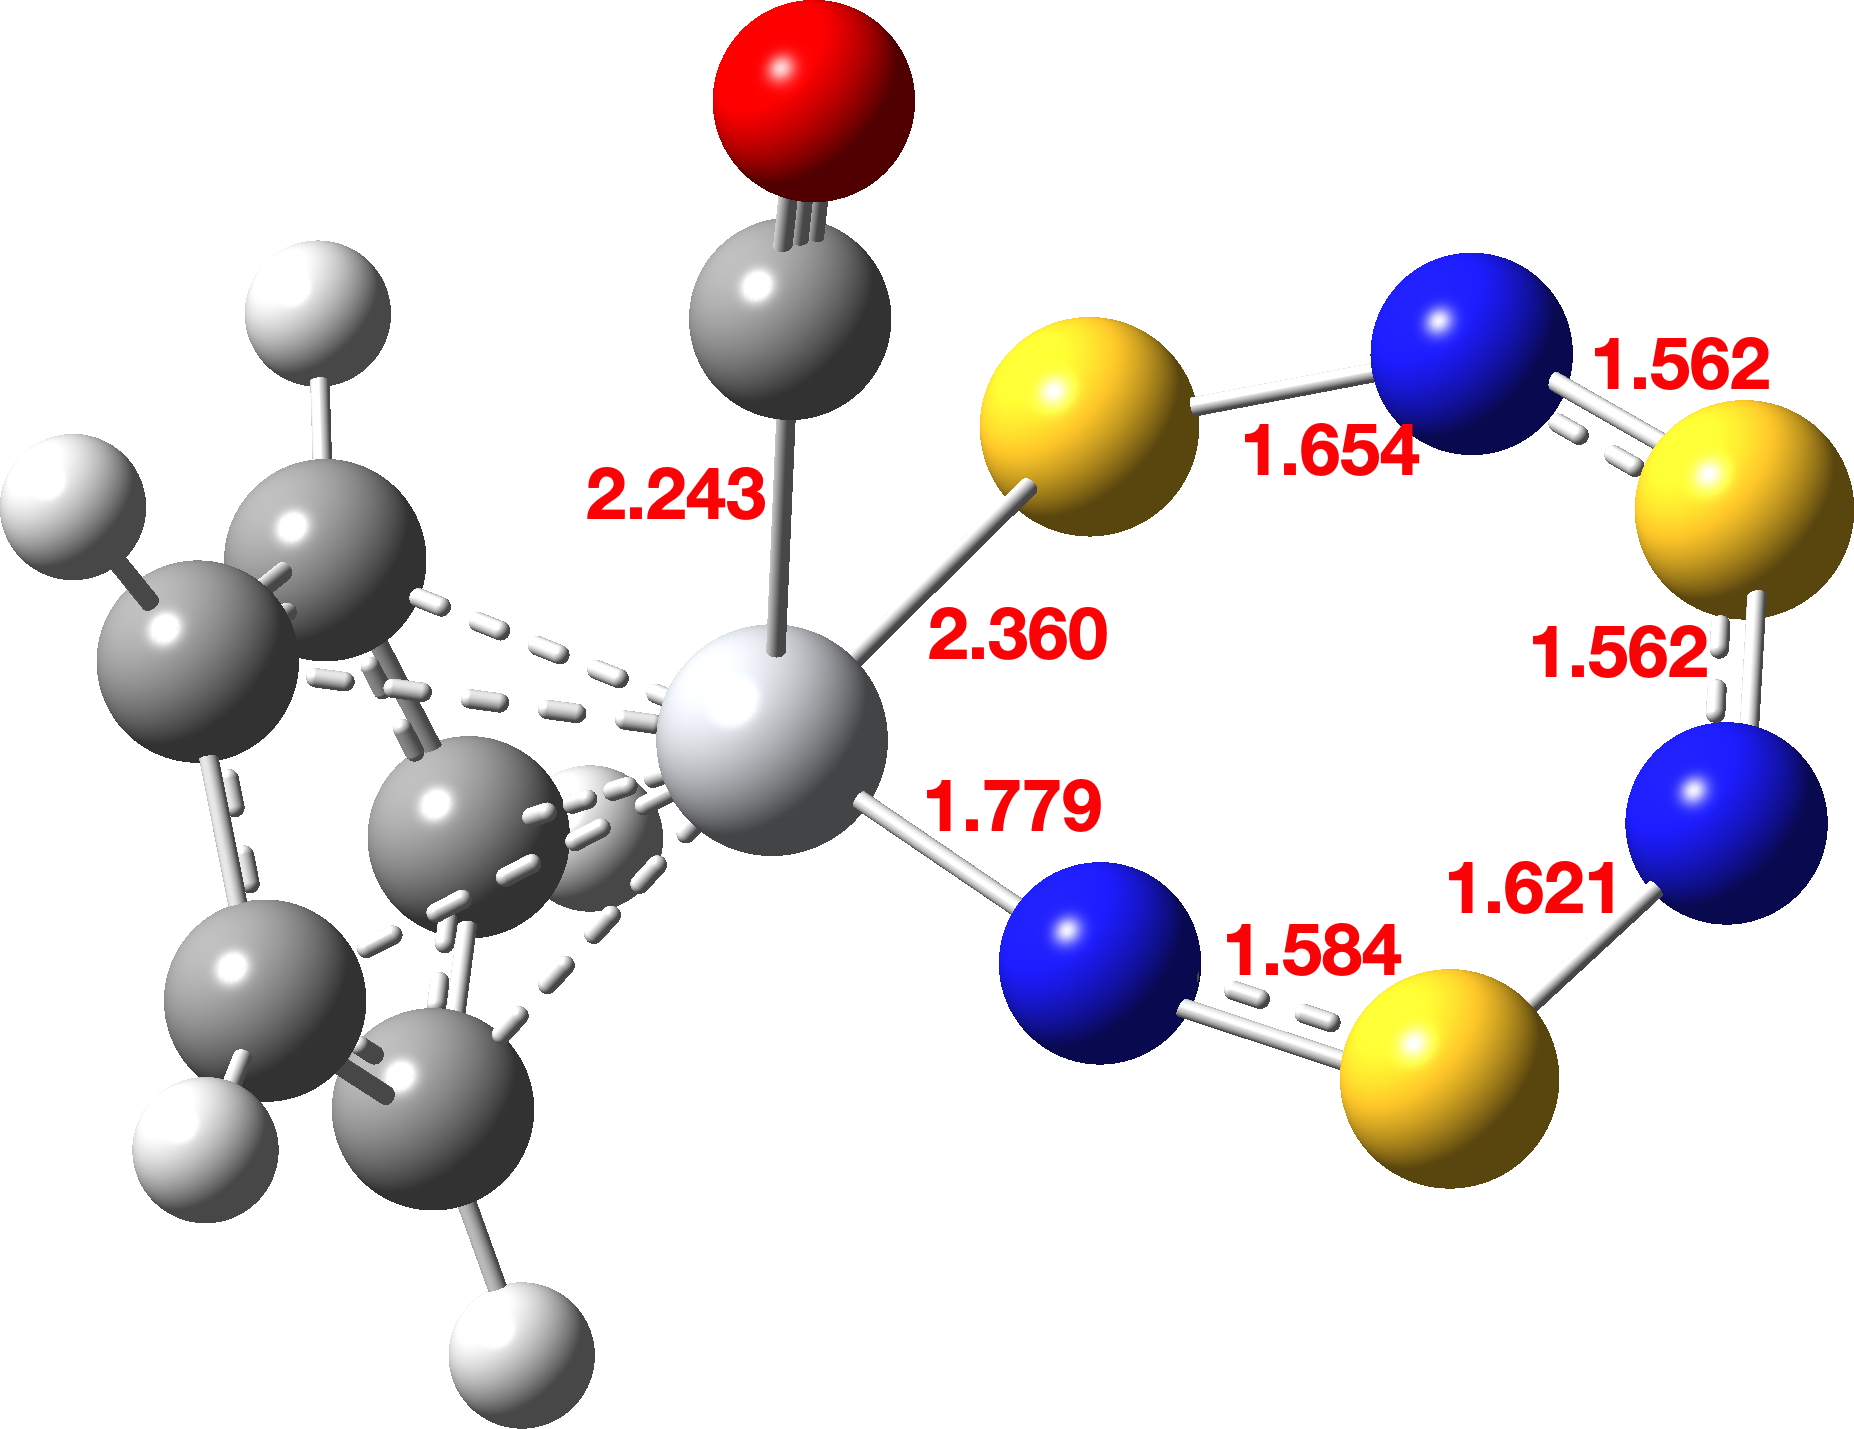
Figure 1. Calculated MN15L/Def2-TZVPP/SCRF=DCM structure of compound 7c.
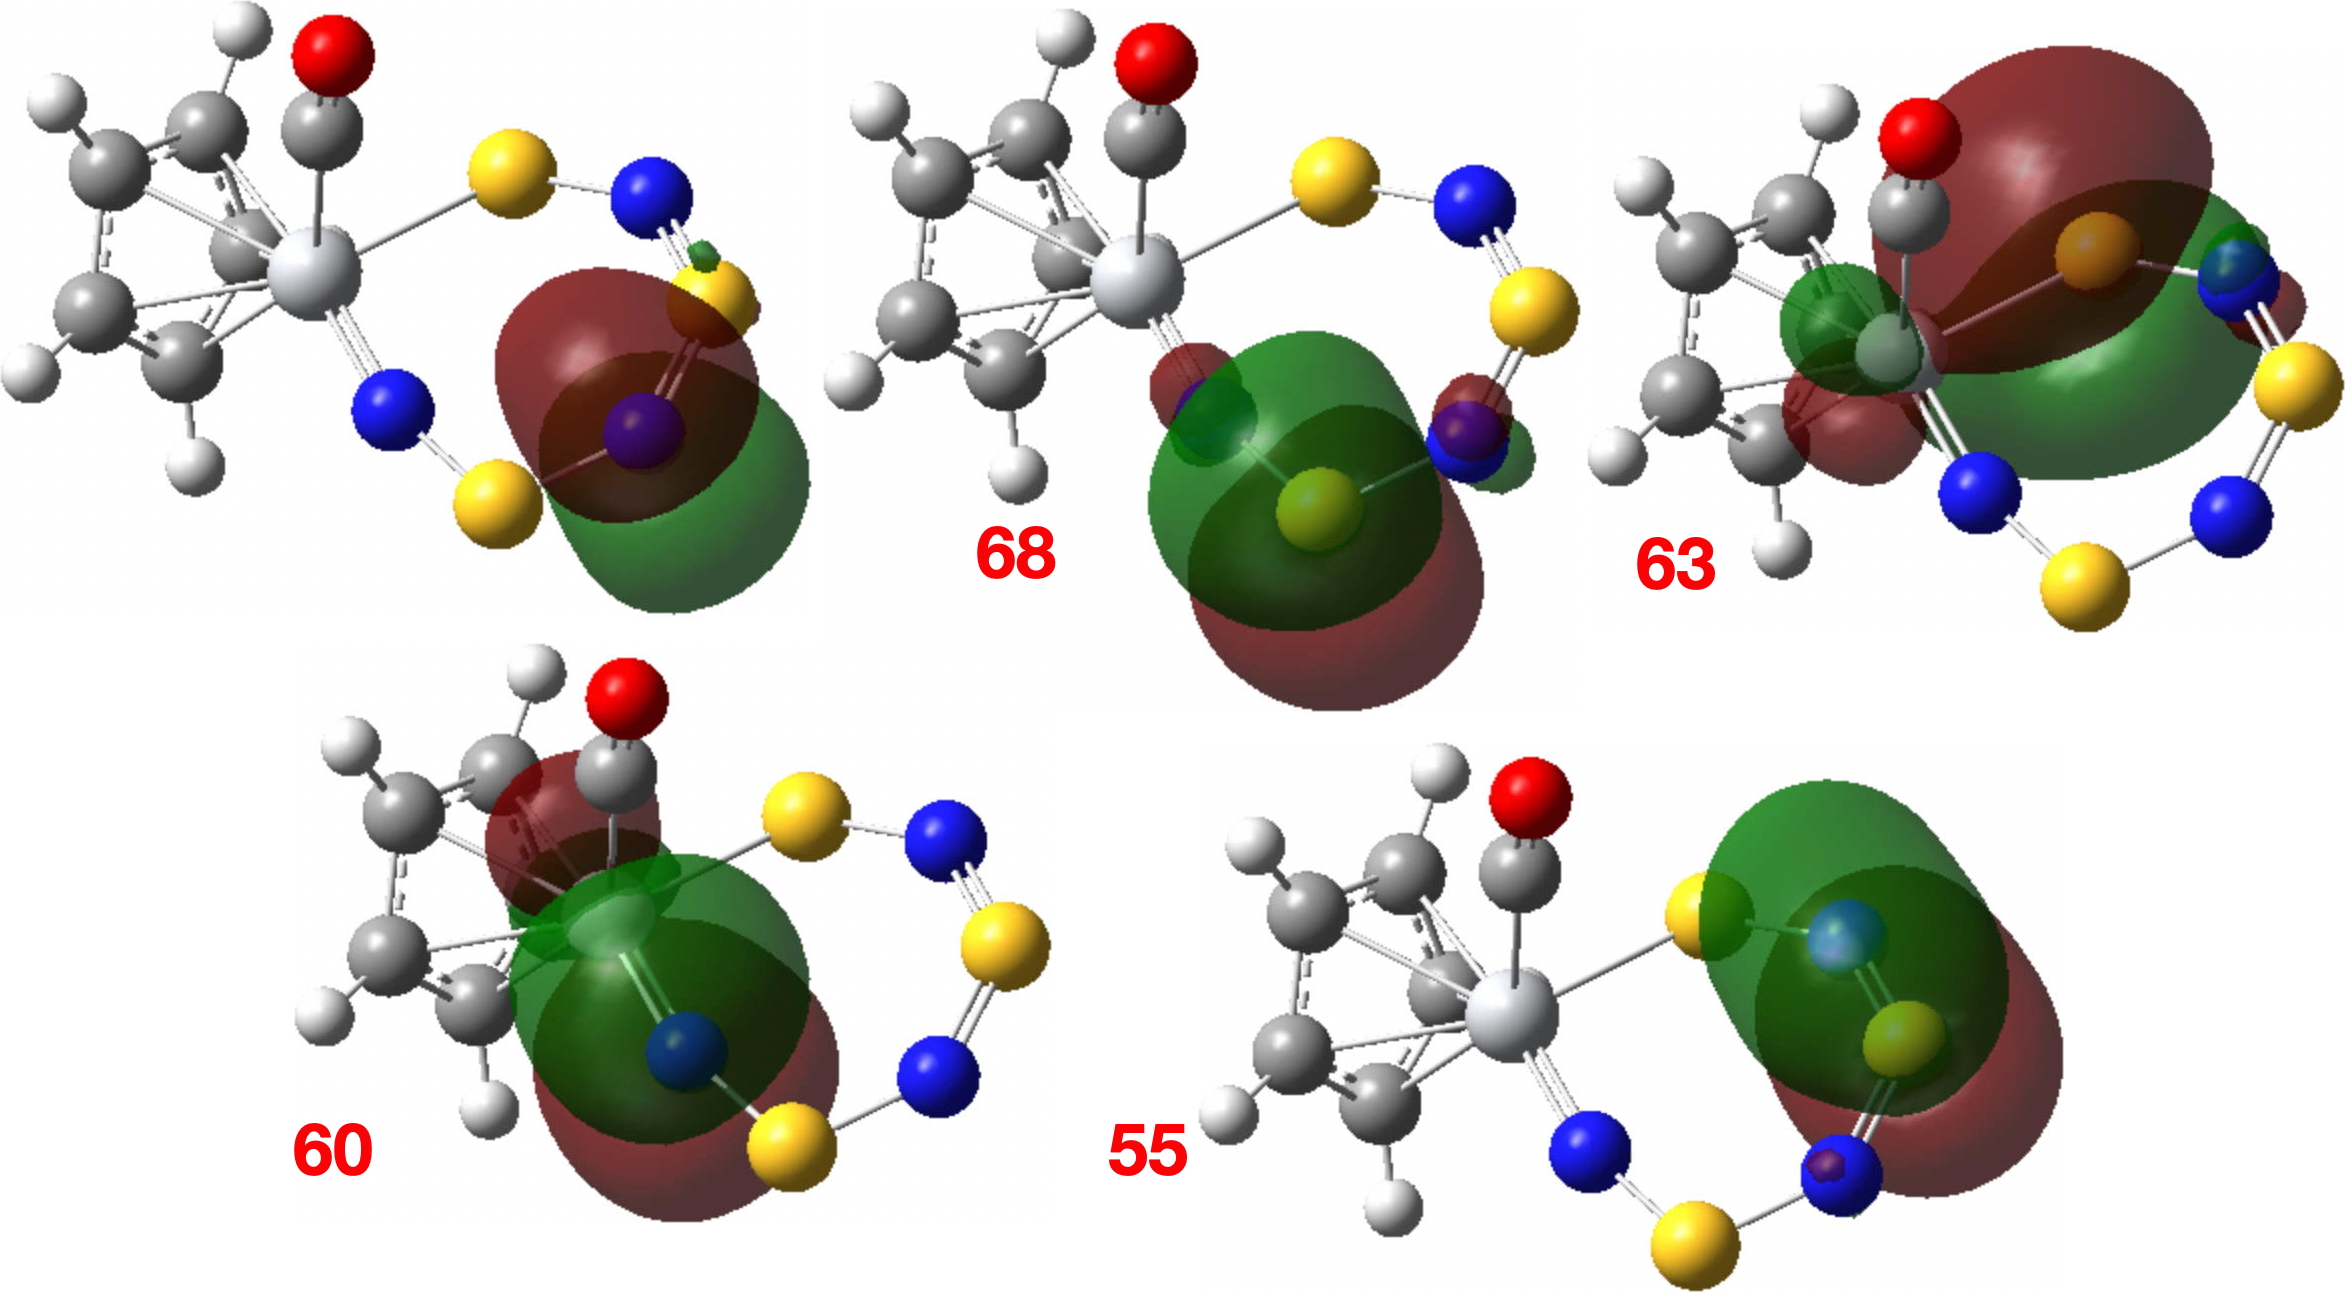
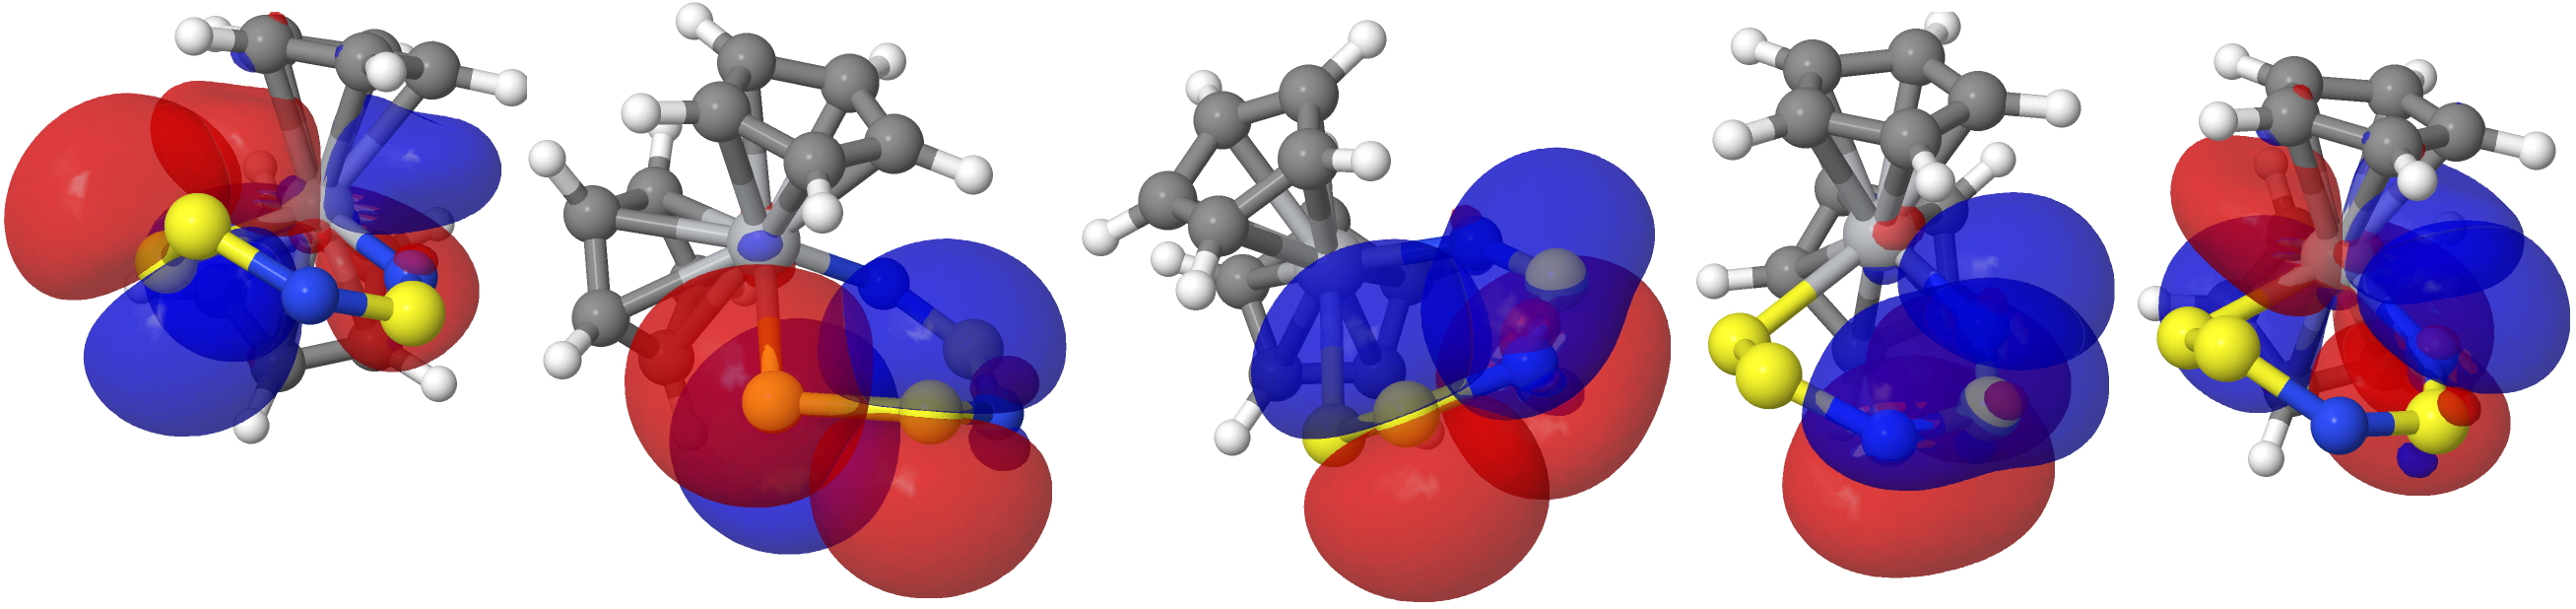
Figure 2. Five NBO7 localised orbitals representing 7c.
A NICS(0) NMR estimate of the aromaticity for 7c gives an antiaromatic value of +9.8 ppm,[20], and hence tending to Möbius antiaromaticity according to the selection rules for 4n+2 (n=2) electrons.[15],[16] This combination of 4n+2 (n=2) conjugated π-electrons and NMR-based antiaromaticity allows one to infer that there must be one orbital phase shift in the conjugated π-electrons, presumably at either the Ti=S centre or the Ti=N centre but not both. This pair of calculations provides an interesting and useful prologue to and calibration of the NICS(0) procedure as an (anti)aromaticity metric in such systems.
Discussion. Molecule 1.
CASZOL10 (molecule 1, scheme 1, Figure 3), has an NBO7‡ Lewis localised structure[21] shown as 1a rather than the literature representation 1[8] (Scheme 1, Figure 4). As with 7c, it comprises three π-bonds and two lone pairs (one N, one S). In contrast to 7c, the NICS(0) = -6.7[22] suggests moderate aromatic character, which as a 10 π-electron aromatic must have either no phase shifts or two. It suggests that, as informed by 7c, the Ti=N bond does not contribute any phase shifts. On the basis of these results, we suggest that molecule 1 can be described as a Hückel type (4n+2) moderately aromatic inorganic metallacycle.
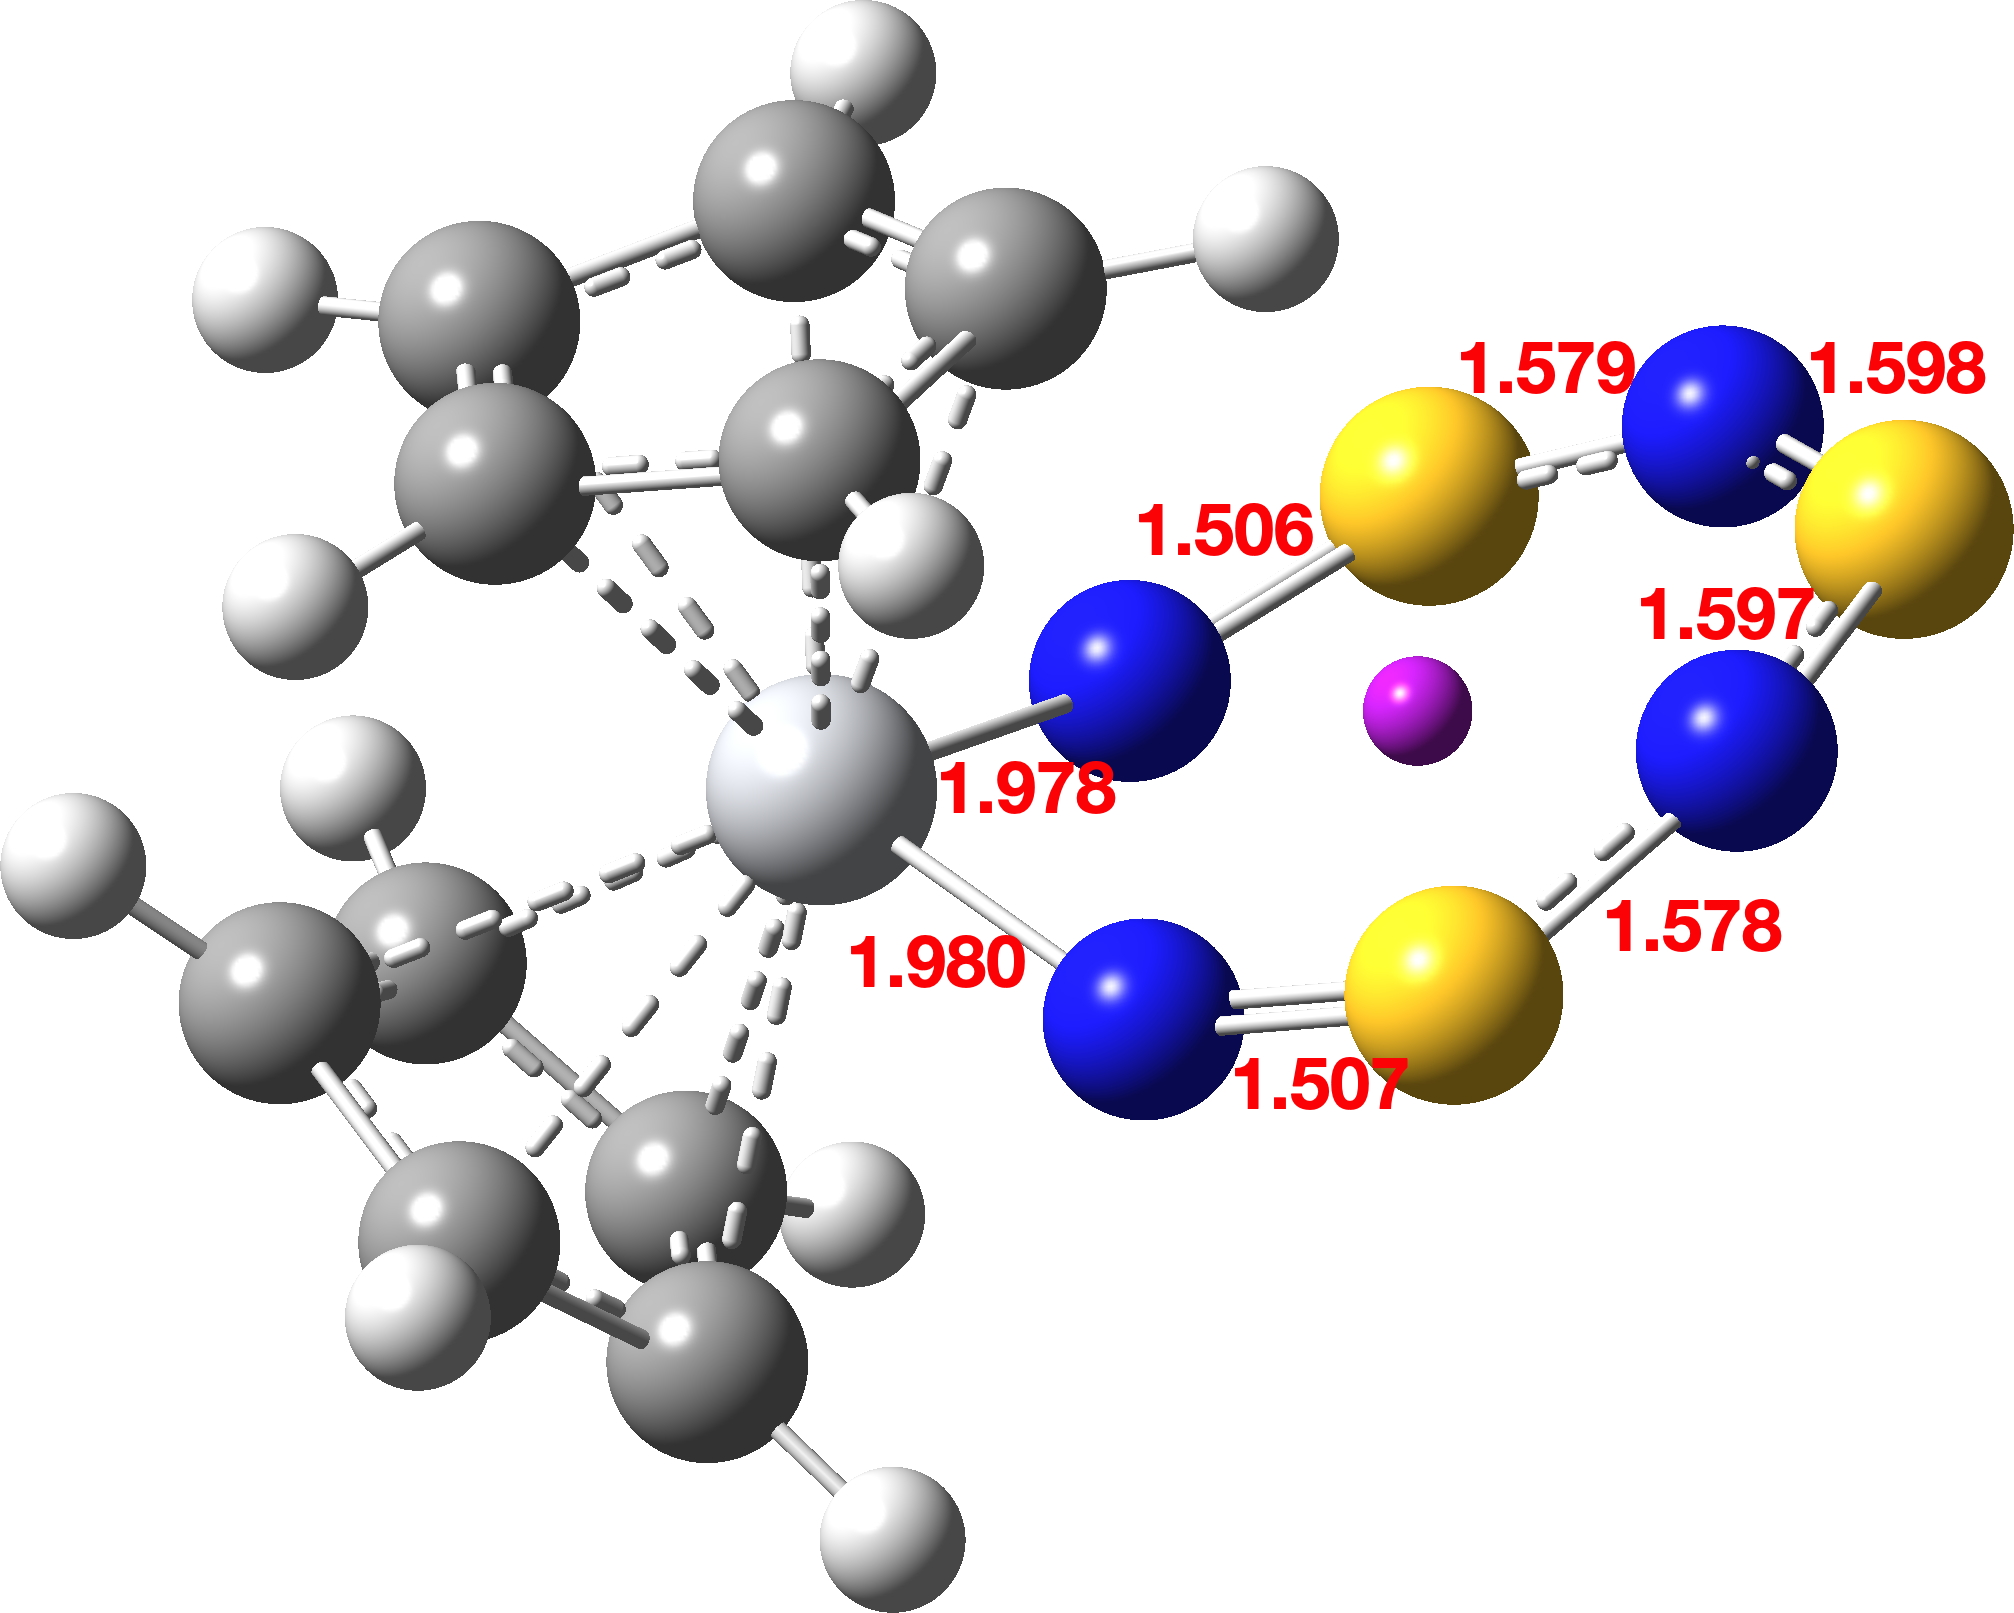
Figure 3. Calculated structure of 1, CASZOL10. The purple point is the NICS probe.
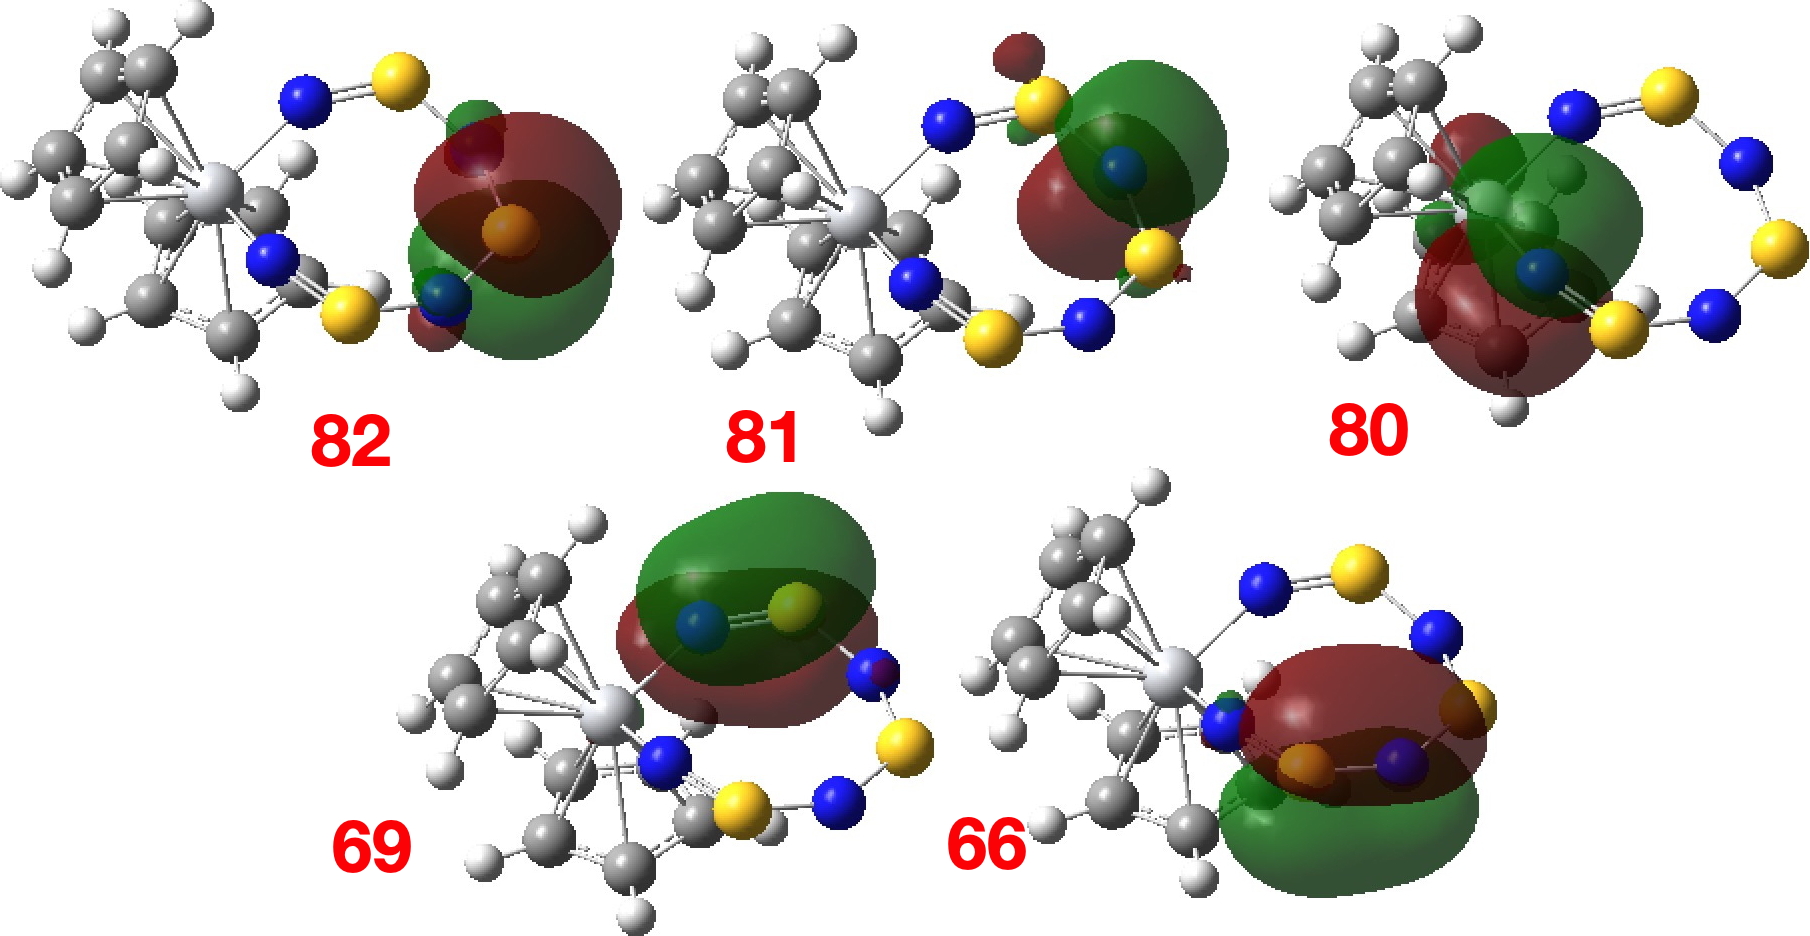
Figure 4. Five NBO7 localised orbitals representing 1a (total NBO7 orbital occupancy 8.39e)
Discussion. Molecule 2.
Molecule 2 (CIWFIX10, Figure 5) has NICS(0) = -6.0 ppm[23] for 8π electrons (one NBO7 double bond and three NBO7 lone pairs[24]) represented as 2a in scheme 1 (Figure 5).
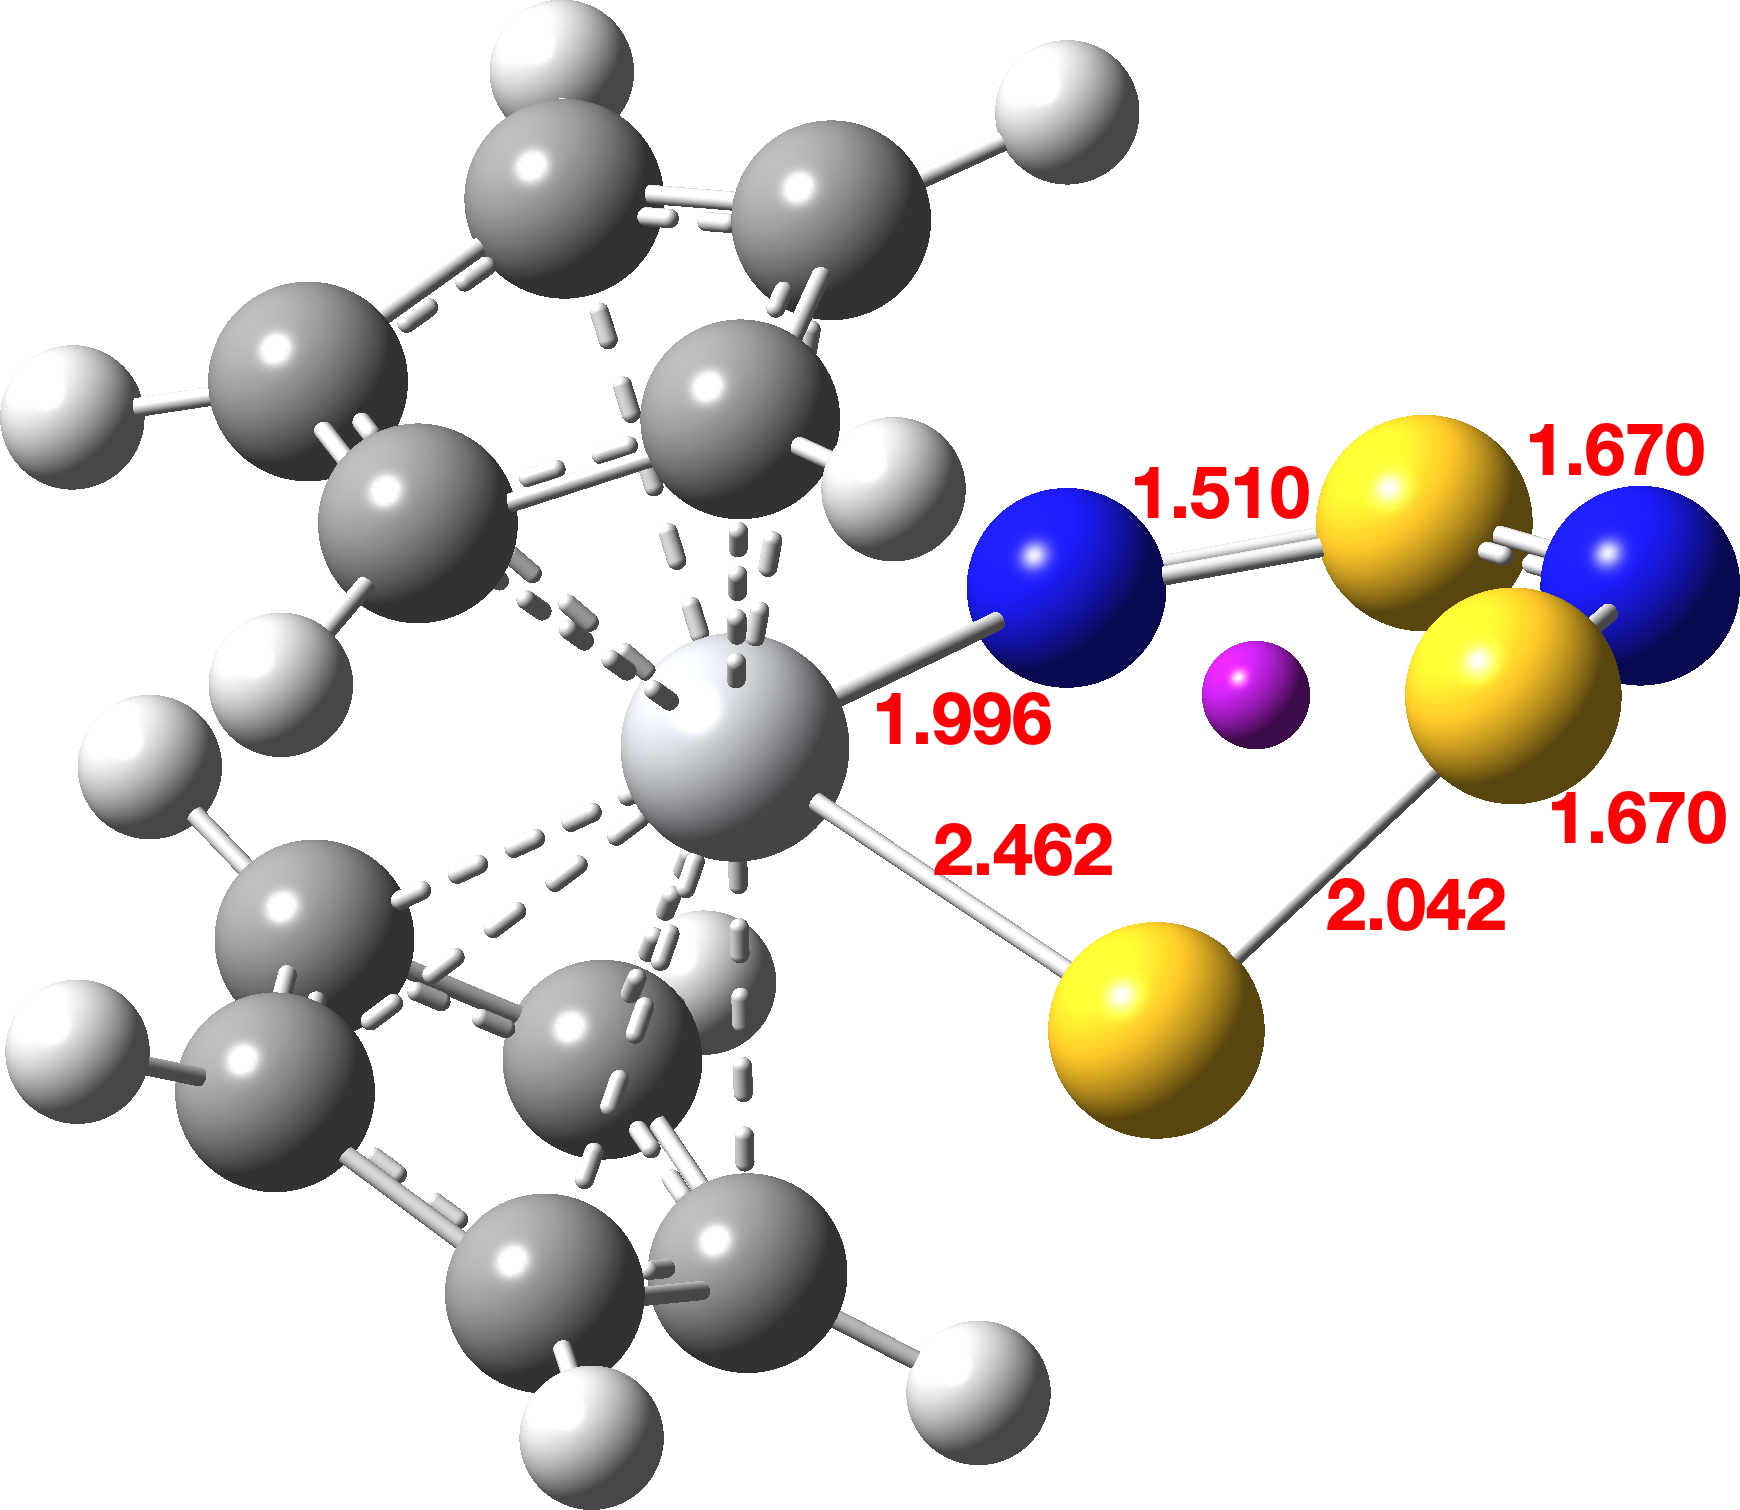
Figure 5. Structure of 2, CIWFIX10. The purple point is the NICS atom.
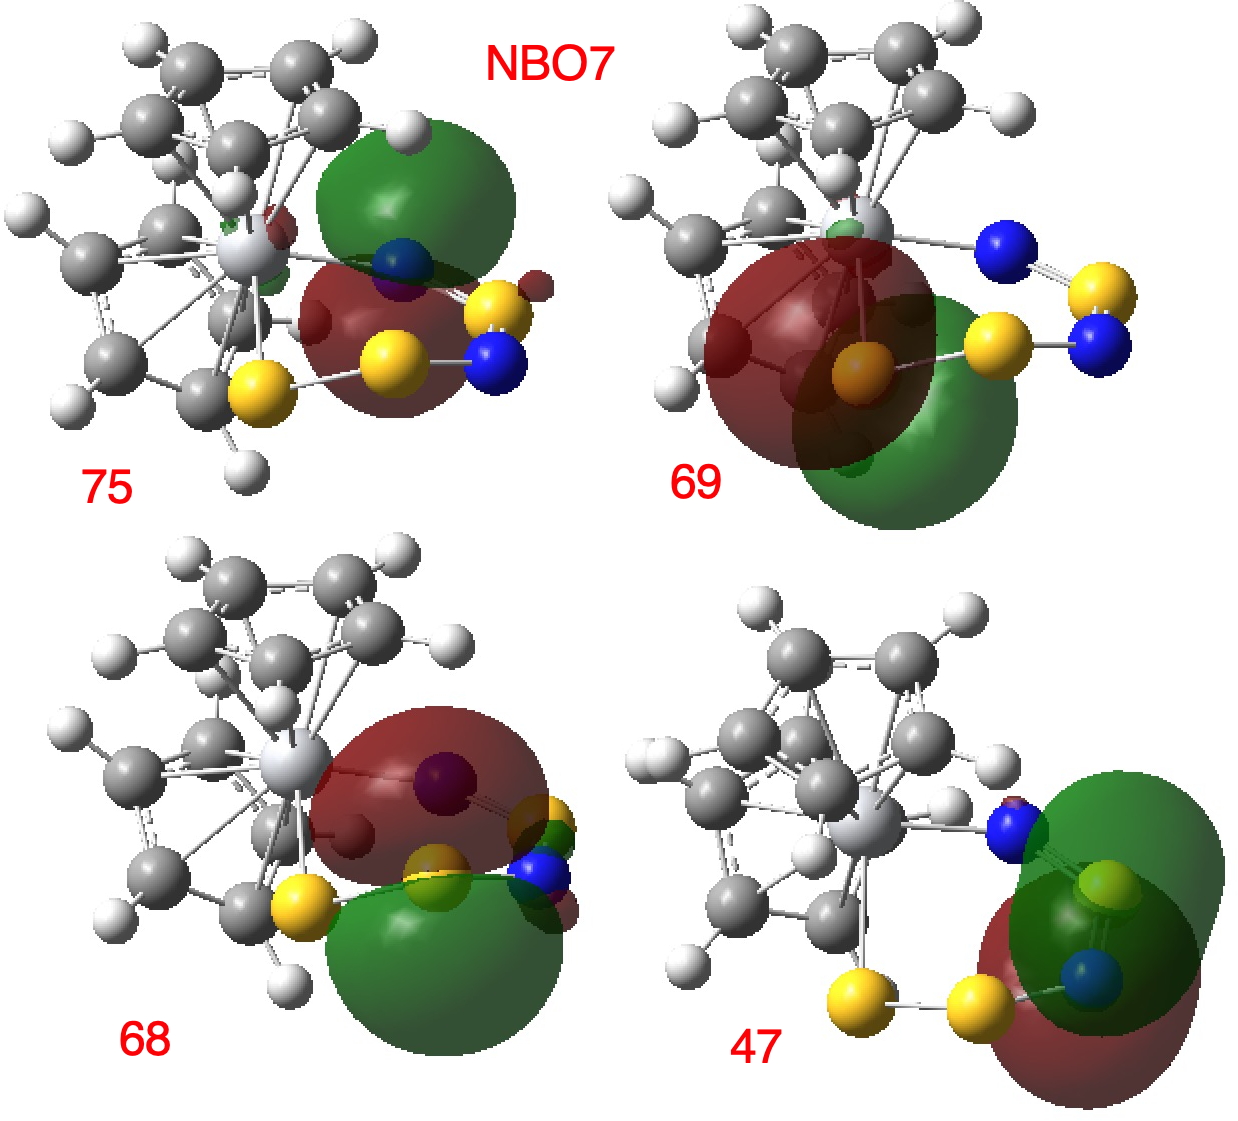
Figure 6. Four NBO7 localised orbitals representing 2b (total NBO orbital occupancy 6.92e).
The moderate NMR-based aromaticity for a 4n (n=2) electron cycle implies Möbius character, [15],[16] whereby significant twisting of the adjacent sulfur-nitrogen ring orbitals allows a phase shift to occur via an unoccupied Ti d-orbital (Figure 7).
Discussion. Unformed Compounds (Scheme3).
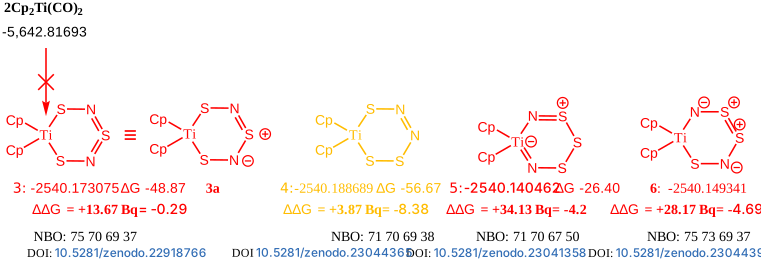
Scheme 3. Unformed compounds 3–6 from the reaction shown in Scheme 1.
Compounds 3-6 are all computed as higher in free energy than 2. Compound 3a (mysteriously not formed in the original report[8]) is +13.7 kcal/mol higher than 2, and clearly non-aromatic (NICS(0) = -0.29 ppm). This may be because the ring is relatively flat and not capable of the twisting required to become Möbius aromatic, something that was possible in 2.[25],[26],[27] Compound 4 (orange in scheme) is interesting since it is only 3.9 kcal/mol higher than 2 and having 8π electrons (one π-bond, three lone pairs) has a NICS(0) of -8.4 ppm[28] appropriate for a Möbius metalla-aromatic and again has a ring capable of the required twisting as per 2. It might be synthesizable if formed by a different method.
Conclusions
Both 1 and 2 are suggested here as examples of inorganic metallaaromatics, respectively of the Hückel and Möbius type,[15],[16] and uniquely with no carbon atoms present in the aromatic ring – a hitherto unrecognised class of aromatic molecule. We here argue that the non-formation of compound 3 as indicated in original article[8] is because it is non-metalla-aromatic, and in its place compound 2 is formed precisely because it IS likely to be metalla-aromatic. It is our expectation that many more such inorganic metalla-aromatics could exist.
‡Use of the older NBO3 procedure is deprecated here, since it is prone to converging for these types of systems to unphysical solutions. We have established that the essential character of the NBO7 orbitals does not depend on the quality of the basis set used[29] or the DFT procedure (albeit tested only for one additional functional, ωB97XD[30],[31]).
This post has DOI: 10.59350/dsyea-mrq85
Authors


References
- https://doi.org/
- R. Jones, J.L. Morris, A.W. Potts, C.W. Rees, D.J. Rigg, H.S. Rzepa, and D.J. Williams, "Electronic and crystallographic structures of trithiadiazepines", Journal of the Chemical Society, Chemical Communications, pp. 398, 1985. https://doi.org/10.1039/c39850000398
- S.T.A.K. Daley, C.W. Rees, and D.J. Williams, "1,3,5,2,4-Trithiadiazepines and 1,3,5,2,4,6-trithiatriazepines, new 10? heteroaromatic systems", Journal of the Chemical Society, Chemical Communications, pp. 55, 1984. https://doi.org/10.1039/c39840000055
- P.N. Jagg, P.F. Kelly, H.S. Rzepa, D.J. Williams, J.D. Woollins, and W. Wylie, "The preparation, X-ray crystal structure and theoretical study of [CoCp <sub>2</sub> ][S <sub>3</sub> N <sub>3</sub> ], (Cp = cyclopentadienyl), a novel stacking compound incorporating multiple C–H ⋯ N(p <sub>π</sub> ) interactions", J. Chem. Soc., Chem. Commun., vol. 0, pp. 942-944, 1991. https://doi.org/10.1039/c39910000942
- A. Haas, and M. Pryka, "New Pathways in Tellurium‐Chalkogen‐Nitrogen Chemistry: Preparations, Structures, and Properties of Telluraheterocycles", Chemische Berichte, vol. 128, pp. 11-22, 1995. https://doi.org/10.1002/cber.19951280103
- J. Galan-Mascaros, A.M. Slawin, J. Derek Woollins, and D.J. Williams, "π-facial interactions between Cl− and [S4N3]+. X-ray crystal structure of [S4N3]Cl", Polyhedron, vol. 15, pp. 4603-4605, 1996. https://doi.org/10.1016/0277-5387(96)00223-9
- T. Chivers, and R.S. Laitinen, "Chalcogen–Nitrogen Chemistry", 2021. https://doi.org/10.1142/12397
- C.G. Marcellus, R.T. Oakley, W.T. Pennington, and A.W. Cordes, "Titanium sulfur nitrogen heterocycles: preparation and molecular structures of titanocene trisulfur tetranitride (.eta.5-C5H5)2TiS3N4 and titanocene trisulfur dinitride (.eta.5-C5H5)2TiS3N2", Organometallics, vol. 5, pp. 1395-1400, 1986. https://doi.org/10.1021/om00138a016
- H. Rzepa, "Cp2TiS3N4, MN15L/Def2-TZVPP G = -2649.576493", 2026. https://doi.org/10.5281/zenodo.22195764
- H. Rzepa, "Cp2TiSSNSN, MN15L/Def2-TZVPP G = -2540.194865, DG = -13.67", 2026. https://doi.org/10.5281/zenodo.22199646
- H. Rzepa, "Cp2TiS3N2, MN15L/Def2-TZVPP G = -2540.173075", 2026. https://doi.org/10.5281/zenodo.22193016
- E.D. Glendening, C.R. Landis, and F. Weinhold, "<i>NBO 7.0</i> : New vistas in localized and delocalized chemical bonding theory", Journal of Computational Chemistry, vol. 40, pp. 2234-2241, 2019. https://doi.org/10.1002/jcc.25873
- H. Rzepa, "Reees 8 (MOs) NBO", 2026. https://doi.org/10.5281/zenodo.22874369
- H. Rzepa, "Reees 8 (MOs) NMR Bq", 2026. https://doi.org/10.5281/zenodo.22876817
- H.S. Rzepa, "Möbius Aromaticity and Delocalization", Chemical Reviews, vol. 105, pp. 3697-3715, 2005. https://doi.org/10.1021/cr030092l
- P.L. Ayers, R.J. Boyd, P. Bultinck, M. Caffarel, R. Carbó-Dorca, M. Causá, J. Cioslowski, J. Contreras-Garcia, D.L. Cooper, P. Coppens, C. Gatti, S. Grabowsky, P. Lazzeretti, P. Macchi, Ã. Martín Pendás, P.L. Popelier, K. Ruedenberg, H. Rzepa, A. Savin, A. Sax, W.E. Schwarz, S. Shahbazian, B. Silvi, M. Solà, and V. Tsirelson, "Six questions on topology in theoretical chemistry", Computational and Theoretical Chemistry, vol. 1053, pp. 2-16, 2015. https://doi.org/10.1016/j.comptc.2014.09.028
- H. Rzepa, "Exploring the effect that causes ring-size specificity of transition metals for polysulfide dianions: Cyclopentadienyl-2,6-di-isopropylphenoxy Titanium pentasulfide.", 2026. https://doi.org/10.59350/6hj5w-5w040
- H. Rzepa, "CpCOTiS3N3 (from Rees system)", 2026. https://doi.org/10.5281/zenodo.22869370
- H. Rzepa, "CpCOTiS3N3 (from Rees system) Compound 7c NBO7", 2026. https://doi.org/10.5281/zenodo.23061061
- H. Rzepa, "CpCOTiS3N3 (from Rees system) NMR Bq Isotropic = -9.8464", 2026. https://doi.org/10.5281/zenodo.22869771
- H. Rzepa, "Cp2TiS3N4, MN15L/Def2-TZVPP Compound 1b, G = -2649.576493 NBO7", 2026. https://doi.org/10.5281/zenodo.22934627
- H. Rzepa, "Cp2TiS3N4, MN15L/Def2-TZVPP G = -2649.576493 NMR Bq Isotropic = 6.6460", 2026. https://doi.org/10.5281/zenodo.22232235
- H. Rzepa, "Cp2TiSSNSN, MN15L/Def2-TZVPP G = -2540.194865, DG = -13.67 NMR Bq Isotropic = 6.0085", 2026. https://doi.org/10.5281/zenodo.22225961
- H. Rzepa, "Cp2TiSSNSN, MN15L/Def2-TZVPP G = -2540.194865, DG = -13.67 Compound 2 NBO7", 2026. https://doi.org/10.5281/zenodo.23018389
- H.S. Rzepa, "A Double-Twist Möbius-Aromatic Conformation of [14]Annulene", Organic Letters, vol. 7, pp. 4637-4639, 2005. https://doi.org/10.1021/ol0518333
- H.S. Rzepa, "Lemniscular Hexaphyrins as Examples of Aromatic and Antiaromatic Double-Twist Möbius Molecules", Organic Letters, vol. 10, pp. 949-952, 2008. https://doi.org/10.1021/ol703129z
- S.M. Rappaport, and H.S. Rzepa, "Intrinsically Chiral Aromaticity. Rules Incorporating Linking Number, Twist, and Writhe for Higher-Twist Möbius Annulenes", Journal of the American Chemical Society, vol. 130, pp. 7613-7619, 2008. https://doi.org/10.1021/ja710438j
- H. Rzepa, "Cp2TiSNNSS MN15L/Def2-TZVPP, G = -2540.188689 NMR Bq Isotropic = 8.3780", 2026. https://doi.org/10.5281/zenodo.22830870
- H. Rzepa, "Cp2TiS3N4, MN15L/Def2-QZVPP Compound 1b, G = -2649.576493 NBO (not 7)", 2026. https://doi.org/10.5281/zenodo.22933586
- H. Rzepa, "Cp2TiS3N4, wB97XD/Def2-TZVPP Compound 1b, G = -2649.576493 NBO (not 7)", 2026. https://doi.org/10.5281/zenodo.23034010
- H. Rzepa, "Cp2TiS3N4, wB97XD/Def2-TZVPP Compound 1b, G = -2649.576493 NBO7", 2026. https://doi.org/10.5281/zenodo.23039008
I know you say that the Möbius twist occurs at Ti in Fig. 7, but it looks to me like the twist occurs across the S-S bond (second-from-left subfigure). Am I misreading the image?
I did some simulation a year or so back to see if the natural “twist” associated with metal d orbitals would make metallabenzenes a rich source of Möbius aromats. Eventually I gave up, because the metal in my examples would always distort out of plane, and it turns out there’s some papers on this. DOI 10.1021/om0701367 says that the LUMO is non-Möbius but the HOMO is Möbius, and both can be lowered in energy as the metal shifts above the ring plane; the counterforce is the disruption to the σ framework, which is weak for metal-ligand bonds. You can particularly see the nonplanarity happening in (your) Fig. 3.
(Of course there is the separate problem that N-S bonds really aren’t π bonds like in benzene. But I see you’ve already posted about that.)
Thanks Jacob for those useful insights. There is still more work to be done to clarify the Möbius components. There is indeed a twist around the S-S bond, but this appears to be below 90° whilst a phase shift requires 180°. Unless the NBO7 code has features I am not familar with, what is needed is a way of computing the dihedral angles between adjacent individual NBOs to quantify the point in the preceding sentence. I might also add that out of plane distortions of up to around 30° do not eliminate aromaticity from benzenoid aromatics. The hypothesis presented in this post is that molecule 2 is formed (and molecule 4 is not far off) because they have some stabilisation due to (Möbius) aromaticity and molecule 3 is not formed because it lacks such stabilisation. Hopefully others can take this up and elaborate it and perchance find other examples. But this problem is non-trivial and I am pretty sure all the answers are NOT present in the post above. At very least, an interesting but forgotten observation has now been teased out of the literature from 1986 and I suspect there are more of this type remaining to be identified. Hey, why not simply hand the problem over to AI? After all, it appears to be working for mathematics!
I thought the following point might be useful since I am unaware it has ever been articulated in the literature. A Möbius system has a single phase shift (π) in the topology of the basis orbitals, and this requires two complete circuits of that basis to return to a starting point. That phase shift can occur in two distinct ways.
1. The phase shift(s) in the form of twist are fully delocalised around the ring. An example might be [14] annulene, where the twist around any two pairs of dihedrals is <20° (see DOI: 10.1021/ol0518333 or 10.1021/jp902176a or 10.1021/ja710438j ). To add to the complexity, there is another topological property known as writhe that also contributes to the total so-called topological property known as the linking number. For 14-annulene the sum of the local twists is actually π, but another contribution of π to the final linking number comes from the writhe!
2. A single phase shift of π is localised to a single centre, such as a metal, along with any contributions from writhe.
Molecule 2 seems to contain a combination of both local and non-local twist and potentially writhe and clearly still has a mystery as to how to incorporate these aspects into the final description.